


OMIM ID:
Pfeiffer Syndrome
Alternate Names
Defective Genes
Clinical Characteristics
Ocular Features
Patients may have extreme proptosis (95%) secondary to shallow orbits and exposure keratitis (41%) is a risk. Hypertelorism, strabismus, and antimongoloid lid slants are common. More rare signs include anterior chamber anomalies and optic nerve hypoplasia.
Systemic Features
Pfeiffer syndrome has been divided into 3 types, of which cases with types 2 and 3 often die young. Type 1 has the more typical features with midface hypoplasia, broad thumbs and toes, craniosynostosis, and often some degree of syndactyly. Adult patients with type 1 may be only mildly affected with some degree of midface hypoplasia and minor broadening of the first digits. Hearing loss secondary to bony defects is relatively common. Cleft palate is uncommon. Airway malformations especially in the trachea can cause respiratory problems.
Genetics
Inheritance
This is a genetically heterogeneous disorder resulting from mutations in at least 2 genes, FGFR1 (8p11.2-p11.1) and FGFR2 (10q26). The less common cases with the latter mutation are allelic to Apert (101200), Crouzon (123500), and Jackson-Weiss (123150) syndromes. Inheritance is autosomal dominant but some cases are only mildly affected. New mutations exhibit a paternal age effect.
Other forms of craniosynostosis in which mutations in FGFR2 have been found are: Beare-Stevenson Syndrome (123790), and Saethre-Chotzen Syndrome (101400).
Pedigree
Autosomal dominant
Autosomal dominant disorders require only one mutation for the disease to be expressed. Since an affected parent has two chromosomes, only one of which has the mutant gene, parents can expect that half (50%) of their children will receive that one and inherit the disease. It is common for individuals that inherit the mutation, however, to not have evidence of the disease (nonpenetrance).
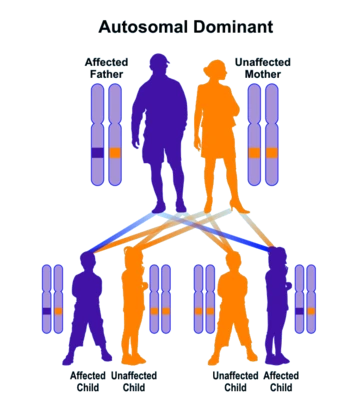
Autosomal dominant inheritance leads to a vertical pattern of transmission