


OMIM ID:
Apert Syndrome
Alternate Names
Defective Genes
Clinical Characteristics
Ocular Features
In 10% of patients, keratitis and corneal scarring occur from the sometimes marked proptosis and corneal exposure. Optic atrophy is present in over 20% of patients. Strabismus, primarily exotropia, is found in more than 70% and various extraocular muscle anomalies may be detectable. Usually the exotropia has a V-pattern with overaction of the inferior oblique muscles while the superior oblique is weak. Amblyopia occurs in nearly 20%. The lid fissures often slant downward and the eyebrows may be interrupted.
Systemic Features
This brachysphenocephalic type of acrocephaly is associated with syndactyly in the hands and feet. Pre- and postaxial polydactyly may be present. There is considerable variation in expression with some patients so mildly affected that they appear virtually normal, whereas others have extreme degrees of brachycephaly with high foreheads, midface hypoplasia, and proptosis secondary to shallow orbits. Imaging often reveals one or more CNS anomalies such as defects of the corpus callosum, partial absence of the septum pellucidum, ventriculomegaly, and sometimes hydrocephalus. A small but significant proportion of patients have some developmental delay and cognitive impairment. Over 39% of patients have a normal IQ.
Genetics
Inheritance
This type of craniosynostosis is caused by mutations in the fibroblast growth factor receptor-2 gene, FGFR2, located at 10q26.13. It is generally considered an autosomal dominant disorder based on familial cases but most occur sporadically. A paternal age effect on mutations has been found. The same gene is mutant in allelic disorders sometimes clinically separated and labeled Crouzon (123500) and Pfeiffer (some cases) (101600) syndromes. Jackson-Weiss syndrome (123150) maps to the same locus. However, this entire group has many overlapping features making classification on clinical grounds alone difficult. Only Apert syndrome is caused by mutations in a single gene whereas other syndromes seem to result from mutations in multiple genes.
Pedigree
Autosomal dominant
Autosomal dominant disorders require only one mutation for the disease to be expressed. Since an affected parent has two chromosomes, only one of which has the mutant gene, parents can expect that half (50%) of their children will receive that one and inherit the disease. It is common for individuals that inherit the mutation, however, to not have evidence of the disease (nonpenetrance).
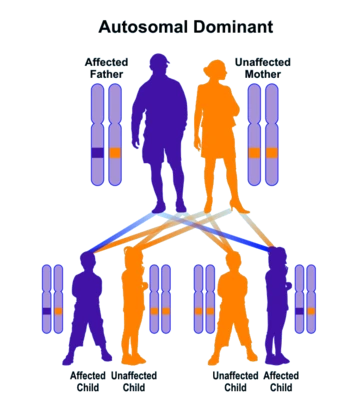
Autosomal dominant inheritance leads to a vertical pattern of transmission