


OMIM ID:
Neurofibromatosis Type II
Alternate Names
Defective Genes
Clinical Characteristics
Ocular Features
It is not uncommon for children to present with visual complaints secondary to cataracts, retinal hamartomas, or optic nerve damage from a schwannoma. The most common eye findings are juvenile cataracts (up to 80% of patients), followed by epiretinal membranes in 12-40%, and retinal tumors in 6-22%. Lens opacities may be located in the posterior subcapsular region or as cortical wedge-shaped opacities in the periphery where they are easily missed if the pupil is not fully dilated. Translucent, grayish epiretinal membranes with white edges are present in a significant number of patients, including children. Elevated retinal hemartomas in the macula are often associated with pigmentary changes. Lisch nodules characteristic of type I neurofibromatosis (162200) are generally not found in type II. Corneal damage may occur as a complication of hypesthesia resulting from damage to the fifth cranial nerve.
CT scans reveal calcification along the optic nerve in a "tram-track" configuration which occurs in 20-30% of patients with NF type II.
Systemic Features
Type II neurofibromatosis often presents in the third or fourth decade of life as hearing loss accompanied by tinnitus and dizziness. A significant proportion of children (30%) present with the same symptoms although they are more likely to complain of visual disturbances. Type II accounts for about 10% of neurofibromatosis cases. Acoustic neurinomas, usually bilateral, are far more common in type II (95%) and are considered diagnostically distinctive by some. Such schwannomas also occur in other cranial and peripheral nerves. Neurofibromas are uncommon but meningiomas, ependymomas, and astrocytomas are seen frequently. Schwannomas can form anywhere along peripheral nerves and at least a third of patients require surgical excision of one or more of these lesions. These account for the majority of skin plaques and lumps and are found in more than half of patients. Café-au-lait spots are uncommon or even absent in many patients with type II. Patients with type II neurofibromatosis do not have the cognitive problems sometimes seen in those with type I.
Longevity overall is reduced. The average patient lives about 15 years after diagnosis and the average age of death is 36 years.
Genetics
Inheritance
Type II neurofibromatosis is an autosomal dominant disorder caused by mutations in the NF2 gene (22q12.2) which encodes neurofibromin-2, sometimes called merlin or schwannomin. This protein product, like neurofibromin in type I (162200), functions as a tumor suppressor. New mutations are responsible for approximately half of cases.
Cognitive deficits and Lisch nodules on the iris are more commonly found in neurofibromatosis type I (162200) but acoustic neuromas are less common. Type I results from mutations in NF1.
Pedigree
Autosomal dominant
Autosomal dominant disorders require only one mutation for the disease to be expressed. Since an affected parent has two chromosomes, only one of which has the mutant gene, parents can expect that half (50%) of their children will receive that one and inherit the disease. It is common for individuals that inherit the mutation, however, to not have evidence of the disease (nonpenetrance).
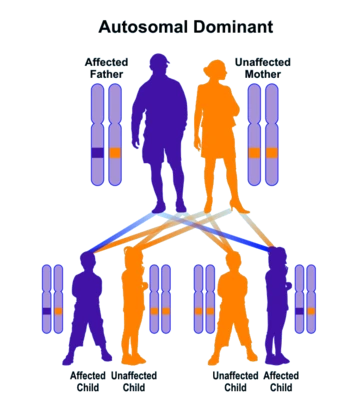
Autosomal dominant inheritance leads to a vertical pattern of transmission