


Clinical Characteristics
Ocular Features
The disorder begins in the first year of life with a band of vascularized opacification inside the limbus. Evidence of inflammation is seen in the anterior stroma and the Bowman membrane becomes replaced by fibrovascular tissue. The disease is recurrent and progressive and there is usually asymmetry between the two eyes. Non-penetrance and considerable variation in expression have been reported. Acute episodes are characterized by photophobia, tearing, mucous discharge, and punctate keratitis. The limbal opacification may progress centrally and eventually leads to a reduction in vision. Deficits in visual acuity may lead to deprivation amblyopia and secondary esotropia.
In a 4 generation family, foveal hypoplasia, iris stromal defects, and ectropion uveae were seen in several of the fifteen affected individuals. It has been suggested that this may be a variant of aniridia.
Systemic Features
No systemic disease has been found.
Genetics
Inheritance
This is an autosomal dominant disorder reported in several multigeneration families. Mutations in the PAX6 gene (11p13) seem to be responsible. The same gene is mutant in Gillespie syndrome (206700), aniridia (106210) and Peters anomaly (604229).
Pedigree
Autosomal dominant
Autosomal dominant disorders require only one mutation for the disease to be expressed. Since an affected parent has two chromosomes, only one of which has the mutant gene, parents can expect that half (50%) of their children will receive that one and inherit the disease. It is common for individuals that inherit the mutation, however, to not have evidence of the disease (nonpenetrance).
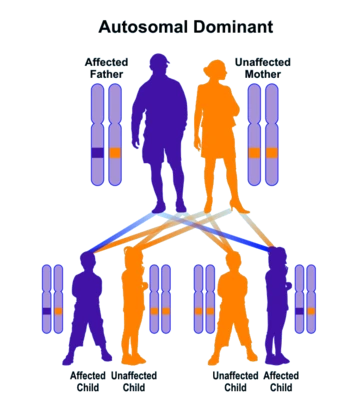
Autosomal dominant inheritance leads to a vertical pattern of transmission