


OMIM ID:
Corneal Dystrophy, Posterior Polymorphous 2
Alternate Names
Defective Genes
Clinical Characteristics
Ocular Features
This is primarily a disease of the posterior cornea although the secondary edema may extend to the epithelium. The disease may be apparent at birth or shortly thereafter by the presence of excrescences or nodules in the endothelial layer with stromal edema. Descemet membrane can be highly irregular in thickness. The endothelial cells in PPCD may acquire some characteristics of epithelial cells.
Systemic Features
No systemic abnormalities have been reported for PPCD2.
Genetics
Inheritance
This is a rare autosomal dominant disorder and few families have been reported. The mutant gene, COL8A2 (1p34.3-p32.3) is the same as that causing early onset Fuchs endothelial dystrophy (136800) and both dystrophies have been described in the same family. The mutation alters the synthesis of alpha 2 chains, part of type VIII collagen, a major component of the Descemet membrance.
For other forms of posterior polymorphous corneal dystrophy, see PPCD3 (609141), and PPPCD1 (122000).
Pedigree
Autosomal dominant
Autosomal dominant disorders require only one mutation for the disease to be expressed. Since an affected parent has two chromosomes, only one of which has the mutant gene, parents can expect that half (50%) of their children will receive that one and inherit the disease. It is common for individuals that inherit the mutation, however, to not have evidence of the disease (nonpenetrance).
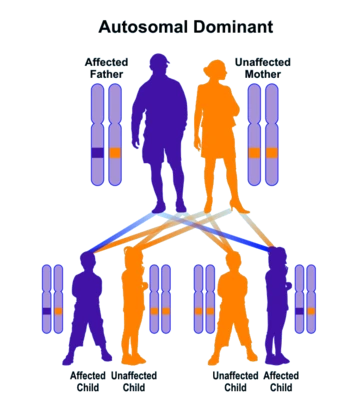
Autosomal dominant inheritance leads to a vertical pattern of transmission