


OMIM ID:
Spinocerebellar Ataxia 42
Alternate Names
Defective Genes
Clinical Characteristics
Ocular Features
Saccadic eye movements with nystagmus and diplopia have been reported (7 of 10 reported patients).
Systemic Features
Cerebellar signs usually have their onset in midlife or later with slow progression. Most patients are mildly to moderately disabled. Dysarthria, dysphagia, and a spastic gait are experienced by the majority of individuals. Hyperreflexia and a positive Babinski sign are commonly presently. Mild cognitive impairment and depression have been seen in a minority of patients.
Brain MRIs show cerebellar hemispheric and vermian atrophy. The cerebral cortex appeared histologically normal in one deceased patient.
Genetics
Inheritance
This disorder is caused by heterozygous mutations in the CACNA1G gene (17q21.33).
Pedigree
Autosomal dominant
Autosomal dominant disorders require only one mutation for the disease to be expressed. Since an affected parent has two chromosomes, only one of which has the mutant gene, parents can expect that half (50%) of their children will receive that one and inherit the disease. It is common for individuals that inherit the mutation, however, to not have evidence of the disease (nonpenetrance).
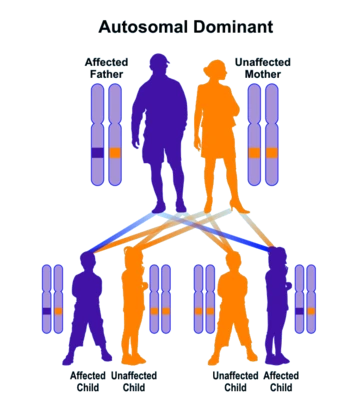
Autosomal dominant inheritance leads to a vertical pattern of transmission