


OMIM ID:
Nanophthalmos 2
Alternate Names
Defective Genes
Clinical Characteristics
Ocular Features
In this condition the axial length of the globe is often only 14-16 mm (normal >20 mm) resulting in extreme hyperopia of +8-25 diopters. Corrected vision is usually 20/40 to 20/80 but 20/200 is not uncommon. The choroid and sclera are thickened in nanophthalmos to a greater degree than seen in common mild hyperopia. While all ocular structures are small in microphthalmia, in nanophthalmos the lens dimensions are generally normal. In a small globe this causes ‘crowding’ of the anterior chamber angles and angle closure glaucoma is a major risk.
Folds in the choroid and retina are common. Choroidal effusions, retinal edema and retinal detachments are not uncommon. The retinal pigment epithelial may have mild window defects. Hypoplasia, cysts, yellowish discoloration, and horizontal striae of the macula have been reported. The foveal reflex is frequently absent corresponding to the lack of a normal foveal pit as revealed by OCT. The foveal avascular zone may be small or absent. The disks often appear crowded. ERGs and VEPs are usually normal. Scleral collagen is abnormal and thickened, leading to the postulation that this interferes with suprachoroidal drainage resulting in effusion and non-rhegmatogenous retinal detachments.
Systemic Features
No systemic disease has been consistently associated with simple nanophthalmos. Individuals with Kenny’s syndrome, Hallerman-Streiff-Francois (234100) syndrome and oculodentodigital dysplasia syndrome (164200) with nanophthalmos have been reported.
Genetics
Inheritance
Nanophthalmos may result from several mutations. Most cases occur sporadically but familial cases suggesting autosomal recessive inheritance (NNO2, 609549) have been reported. The mutation is a frameshift insertion, 1143C, in the MFRP gene on chromosome 11 (11q23.3) and has been found in the homozygous configuration in several families. The protein product has a domain that may be related to the Frizzled family of transmembrane cell-cell signaling molecules responsible for regulation of growth and differentiation. In this connection, it is of interest that this gene is highly expressed in the retinal pigment epithelium.
It seems that at least two dominant mutations can also cause nanophthalmos. One (NNO3, 611897), located on chromosome 2 (2q11-q14), has been identified in a large Chinese pedigree although the molecular mutation remains unknown. Another, NNO1, (600165), has also been mapped to chromosome 11 but at 11p. The molecular mutations also remain unknown.
Homozygous mutations in serine protease PR2258 have also been reported in several families with nanophthalmos.
Pedigree
Autosomal dominant
Autosomal dominant disorders require only one mutation for the disease to be expressed. Since an affected parent has two chromosomes, only one of which has the mutant gene, parents can expect that half (50%) of their children will receive that one and inherit the disease. It is common for individuals that inherit the mutation, however, to not have evidence of the disease (nonpenetrance).
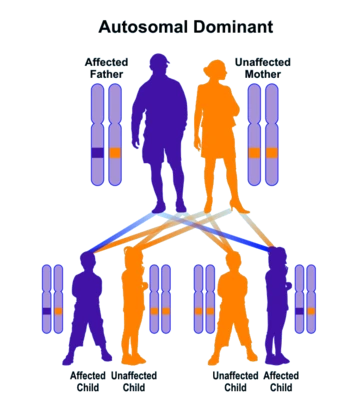
Autosomal dominant inheritance leads to a vertical pattern of transmission
Autosomal recessive
In order for autosomal recessive disorders to be expressed, offspring generally must inherit two mutations, one from each carrier parent. Carriers with only one mutation, such as the parents, do not have clinical disease. Note that carrier parents can expect that 1 in 4 children (25%) will inherit both mutations and have the disorder, 2 in 4 children (50%) will be carriers like their parents, while 1 in 4 children (25%) inherit neither mutation.

In order for autosomal recessive disorders to be expressed, offspring generally must inherit two mutations, one from each carrier parent. Carriers with only one mutation, such as the parents, do not have clinical disease. Note that carrier parents can expect that 1 in 4 children (25%) will inherit both mutations and have the disorder, 2 in 4 children (50%) will be carriers like their parents, while 1 in 4 children (25%) inherit neither mutation.