


OMIM ID:
Foveal Hypoplasia 1
Alternate Names
Defective Genes
Clinical Characteristics
Ocular Features
This is a poorly defined syndrome with features overlapping aniridia, hereditary keratitis, ocular albinism, and iris anomalies as in Peters anomaly. However, presenile cataracts seem to be unique to this disorder. The foveal hypoplasia may occur without other anomalies although the fundus is usually lightly pigmented. As expected, acuity is subnormal from birth, in the range of 20/50, and dyschromatopsia may be present. Some patients have nystagmus. Weak iris transillumination has been reported and a small limbal pannus may be present. Lens opacities may become visually significant in the third to fourth decade of life. OCT has shown abnormal foveal thickness with multiple inner retinal layers somewhat similar to the situation in oculocutaneous albinism (203100) and it has been suggested that 'foveal dysplasia' is a better description than 'foveal hypoplasia'.
Systemic Features
No systemic disease is present.
Genetics
Inheritance
This disorder is associated with mutations in the PAX6 gene (11p13) and inherited as an autosomal dominant.
The protein product of the PAX6 gene is a transcription factor that attaches to DNA and regulates the expression of other genes. PAX6 plays a major role primarily in development of the eye and central nervous system but evidence suggests it is also active postnatally. Hundreds of mutations have been found in disorders such as hereditary keratitis, aniridia, Peters anomaly, hypoplasia and colobomas of the optic nerve. This database contains 8 conditions in which mutations in PAX6 seem to be responsible, including syndromal conditions such as Stromme and Gillespie syndromes in which there may be cognitive disabilities.
True isolated foveal hypoplasia without lens or corneal disease does exist as well but this condition (FVH2) is not well defined. Homozygous mutations in SLC38A8 have been found to cosegregate with this form of foveal hypoplasia among families of Jewish Indian ancestry. Hypopigmentation is not a feature of isolated foveal hypoplasia secondary to such mutations but misrouting of optic nerve axons may be present. Nystagmus and reduced vision but no anterior segment abnormalities were present.
With the widespread utilization of OCT measurements, we have learned that underdevelopment of the fovea can be a feature of numerous ocular disorders (more than 20 in this database). In most conditions, the foveal dysplasia is part of a disease complex as in foveal hypoplasia with anterior segment dysgenesis (609218).
Pedigree
Autosomal dominant
Autosomal dominant disorders require only one mutation for the disease to be expressed. Since an affected parent has two chromosomes, only one of which has the mutant gene, parents can expect that half (50%) of their children will receive that one and inherit the disease. It is common for individuals that inherit the mutation, however, to not have evidence of the disease (nonpenetrance).
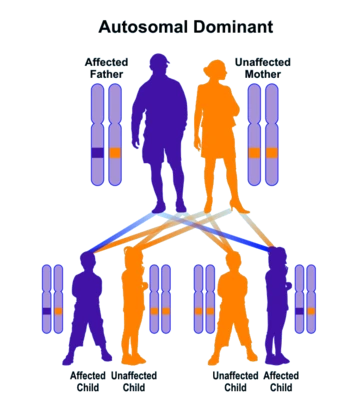
Autosomal dominant inheritance leads to a vertical pattern of transmission