


Clinical Characteristics
Ocular Features
Many patients have few ocular findings beyond the usual synophyrs, a highly arched brow with hypertrichosis, and long eyelashes. Synophrys is often prominent. However, some also have significant ptosis, nystagmus, and high refractive errors. Optic pallor and a poor macular reflex have also been reported.
Systemic Features
The facial features may be distinctive with low anterior hairline, anteverted nares, maxillary prognathism, long philtrum, crescent-shaped mouth and, of course, the bushy eyebrows and long lashes (in 98%). Mental and growth retardation are common while many patients have features of the autism spectrum and tend to avoid social interactions. The lips appear thin, the mouth is crescent-shaped, the head is often small, the teeth are widely spaced, and the ears are low-set. The hands are often deformed with a proximally positioned thumb and metacarpophalangeal deformities. It is stated that the middle phalanx of the index finger is always hypoplastic. Other limb abnormalities of both upper (95%) and lower extremities are common. Urinary tract abnormalities have been found in 41% of patients. Middle ear effusions often lead to conductive hearing loss but 80% of patients have a sensorineural hearing deficit.
Genetics
Inheritance
This disorder is caused by mutations in genes encoding components of the cohesion complex. Most cases occur sporadically but numerous familial cases suggest autosomal dominant inheritance. However, since at least three genes code for components of the cohesion complex including one located on the X-chromosome (610759), familial cases reported earlier without genotyping have created some confusion. Hence, even autosomal recessive inheritance has been suggested in some families. Genetic counseling should be family-specific based on the genotype and family pattern.
About 50% of cases result from mutations in the NIPBL gene (122470; 5p13.1) but less than 1% have an affected parent and the recurrence risk for sibs is similar. The X-linked form of CDLS (300590; Xp11.22-p11.21) is caused by a mutation in the SMC1A gene, and a mild form (610759) results from mutations in the SMC3 gene (10q25). Mutations in RAD21 (8q24) have been found in patients with milder disease and atypical presentations (614701).
A CDLS phenotype can also result from a specific duplication of a 3q 26-27 band.
Pedigree
Autosomal dominant
Autosomal dominant disorders require only one mutation for the disease to be expressed. Since an affected parent has two chromosomes, only one of which has the mutant gene, parents can expect that half (50%) of their children will receive that one and inherit the disease. It is common for individuals that inherit the mutation, however, to not have evidence of the disease (nonpenetrance).
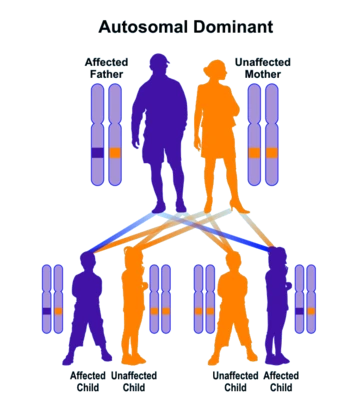
Autosomal dominant inheritance leads to a vertical pattern of transmission
Autosomal recessive
In order for autosomal recessive disorders to be expressed, offspring generally must inherit two mutations, one from each carrier parent. Carriers with only one mutation, such as the parents, do not have clinical disease. Note that carrier parents can expect that 1 in 4 children (25%) will inherit both mutations and have the disorder, 2 in 4 children (50%) will be carriers like their parents, while 1 in 4 children (25%) inherit neither mutation.

In order for autosomal recessive disorders to be expressed, offspring generally must inherit two mutations, one from each carrier parent. Carriers with only one mutation, such as the parents, do not have clinical disease. Note that carrier parents can expect that 1 in 4 children (25%) will inherit both mutations and have the disorder, 2 in 4 children (50%) will be carriers like their parents, while 1 in 4 children (25%) inherit neither mutation.