


OMIM ID:
Corneal Dystrophy, Fuchs Endothelial, Late Onset 2
Alternate Names
Defective Genes
Clinical Characteristics
Ocular Features
The signs and symptoms of this disorder are similar to those of other adult endothelial dystrophies. Guttae appear in the fourth or fifth decade of life and gradually increase in number. Diffuse corneal edema eventually develops with a corresponding decrease in acuity. In late stages the corneal edema involves all layers including the epithelium, leading to painful corneal erosions.
Systemic Features
Some patients with FECD3 report hearing impairment but this has not been studied and may be simply an age-related association. It is of interest that among an unclassified series of patients with FCD the frequency of hearing loss was higher than in matched controls.
Genetics
Inheritance
A mutation in LOXHD1 (18q21.2-q21.32) was originally thought to be responsible for this form of Fuchs in a multigenerational pedigree but is now considered an insignificant variant. More recent evidence suggests that heterozygous trinucleotide repeat expansions in the TCF4 transcription factor gene at 18q22 are responsible.
There is considerable genetic heterogeneity in adult endothelial dystrophy which makes the nosology confusing especially since the clinical features are similar. A similar late onset autosomal dominant disease [Fuchs Endothelial Dystrophy, Late Onset (610158)], sometimes labeled FCD2, may result from mutations on chromosome 13, or from changes in ZEB1 on chromosome 10. Many cases are sporadic, however, and additional genotyping will be necessary in individuals to further clarify the classification of late-onset Fuchs endothelial dystrophy.
There is also an early onset form of Fuchs endothelial dystrophy, (136800).
Pedigree
Autosomal dominant
Autosomal dominant disorders require only one mutation for the disease to be expressed. Since an affected parent has two chromosomes, only one of which has the mutant gene, parents can expect that half (50%) of their children will receive that one and inherit the disease. It is common for individuals that inherit the mutation, however, to not have evidence of the disease (nonpenetrance).
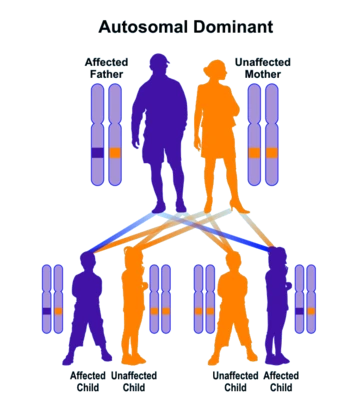
Autosomal dominant inheritance leads to a vertical pattern of transmission